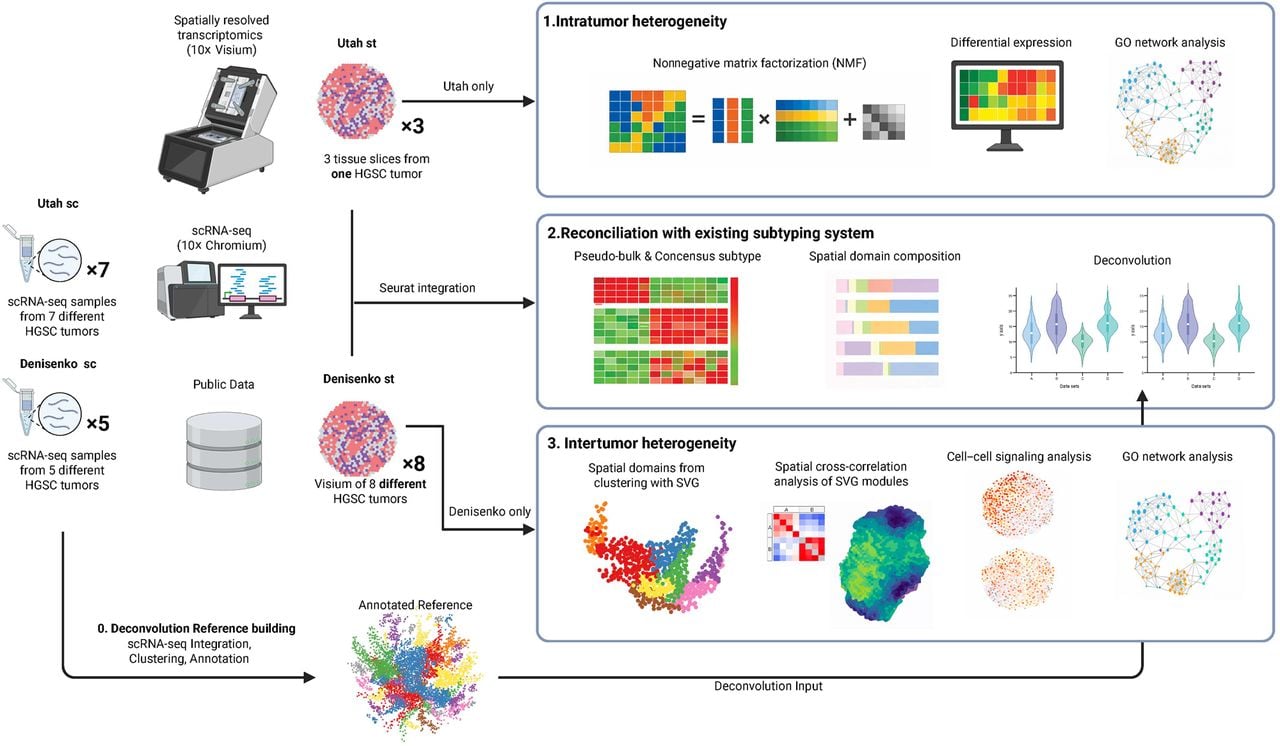
Integrated scRNA-seq and Visium spatial transcriptomics workflow for characterizing heterogeneity in high-grade serous carcinoma (HGSC). Public scRNA-seq and Visium data sets from HGSC tumors reported by Denisenko et al. (2024) are integrated with newly generated scRNA-seq and Visium data to investigate intra- and intertumor heterogeneity. All scRNA-seq data sets are jointly integrated to define cell states and to serve as a reference for downstream spatial transcriptomics deconvolution. Nonnegative matrix factorization (NMF) is applied to spatially variable genes in newly collected Visium samples from a single tumor to identify intratumoral spatial domains. All Visium samples are pseudobulked, consensus-subtyped, and deconvolved to assess the relationship between cell-type composition and established HGSC subtypes. Visium samples from Denisenko et al. are further analyzed to identify spatial domains using spatially variable genes (SVGs) and to perform Gene Ontology (GO) enrichment. Finally, spatial cross-correlation and cell–cell signaling analyses are conducted on a per-tumor basis to characterize intertumor heterogeneity and conserved spatial patterns.